A Form FDA 483 and an FDA Warning Letter are not the same document. A 483 lists the investigator's observations
For medical devices, an FDA adverse event report is a Medical Device Report (MDR) under 21 CFR Part 803. The
A De Novo review is FDA's risk-based classification route for a novel device that has no legally marketed predicate, but
Device listing is the part of FDA registration that most teams touch once a year and otherwise forget. That is
Section 513 of the FD&C Act, added by the Medical Device Amendments of 28 May 1976, sets up FDA's three
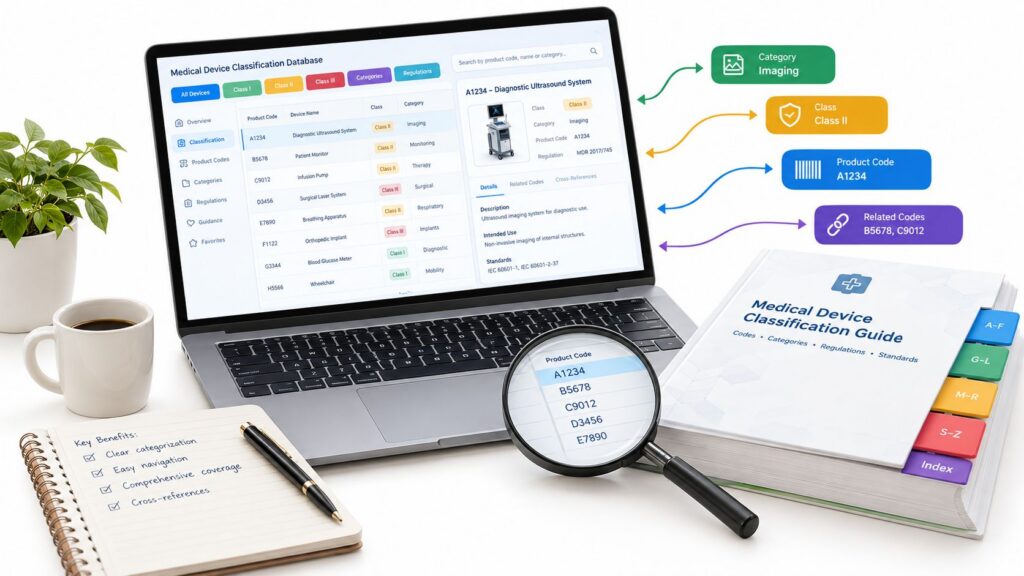
FDA has classified more than 1,700 distinct types of devices and organised them into 16 medical specialty panels in 21
The MDR, Regulation (EU) 2017/745, was adopted on 5 April 2017 and has applied since 26 May 2021, replacing Directives
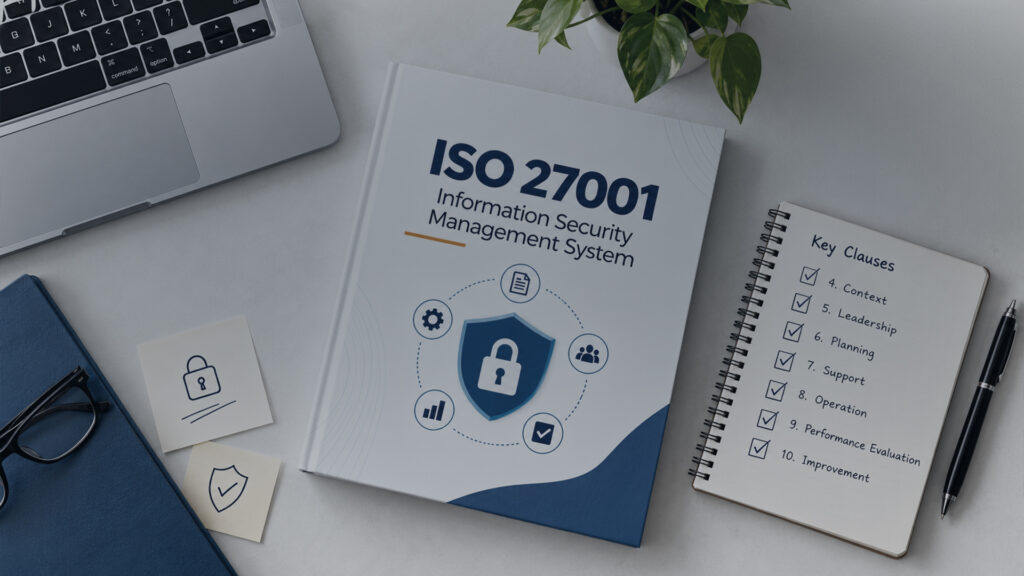
ISO 27001 Requirements: A Clause-by-Clause Reference ISO/IEC 27001 has been the reference standard for information security management systems since 2005,
21 CFR Part 11 began as a permission, not a mandate. The FDA published it in March 1997, in 62
Explore AI-enabled SaMD requirements with our easy step-by-step guide.
Cookies help us improve your experience on our website. By using our site, you consent to the use of cookies as described in this policy.