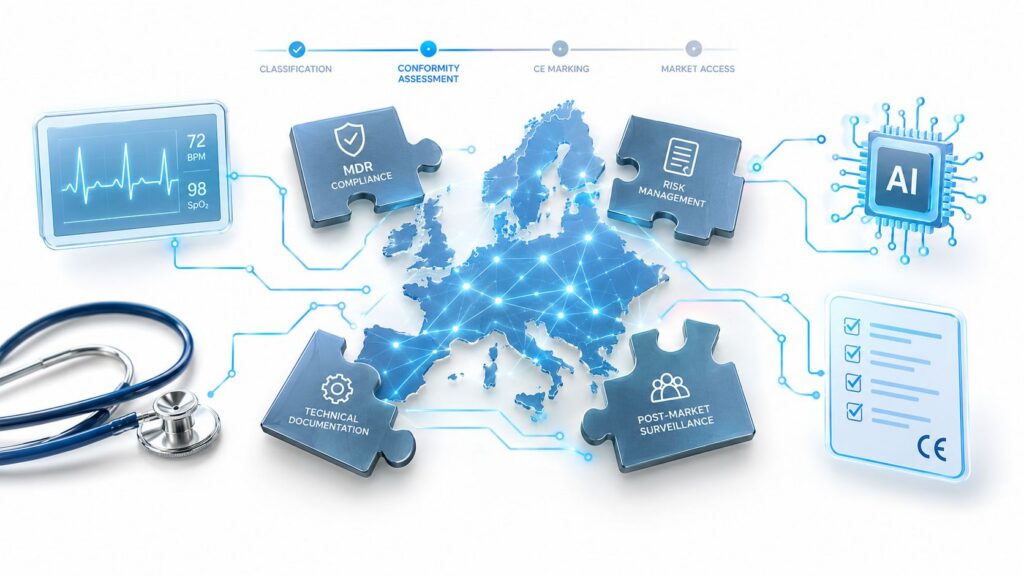
For medical devices, an FDA adverse event report is a Medical Device Report (MDR) under 21 CFR Part 803. The
A De Novo review is FDA's risk-based classification route for a novel device that has no legally marketed predicate, but
Device listing is the part of FDA registration that most teams touch once a year and otherwise forget. That is
ISO 27001 Requirements: A Clause-by-Clause Reference ISO/IEC 27001 has been the reference standard for information security management systems since 2005,
21 CFR Part 11 began as a permission, not a mandate. The FDA published it in March 1997, in 62
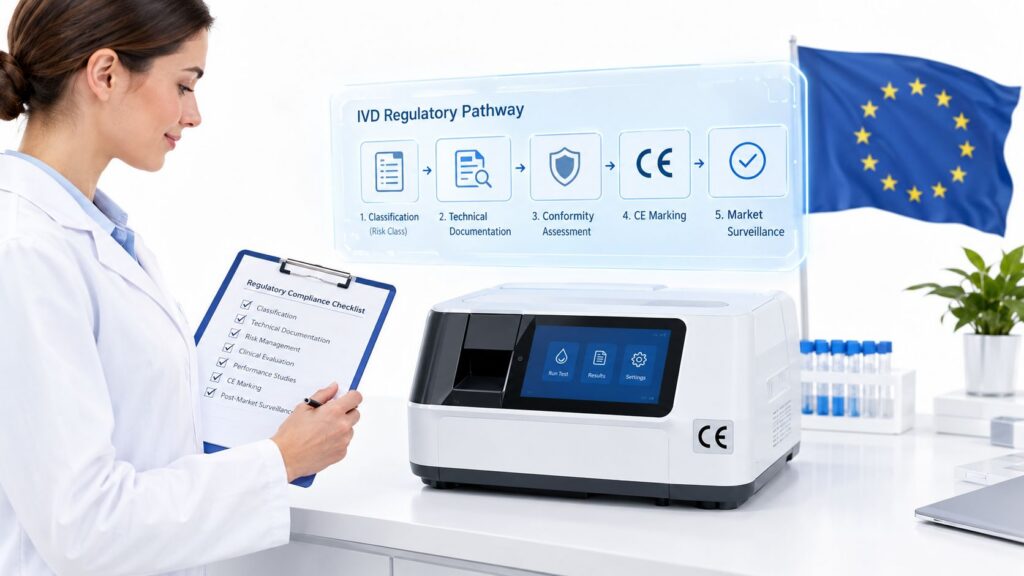
The shift from Directive 98/79/EC to Regulation (EU) 2017/746, the In Vitro Diagnostic Regulation, changed the route to IVD CE
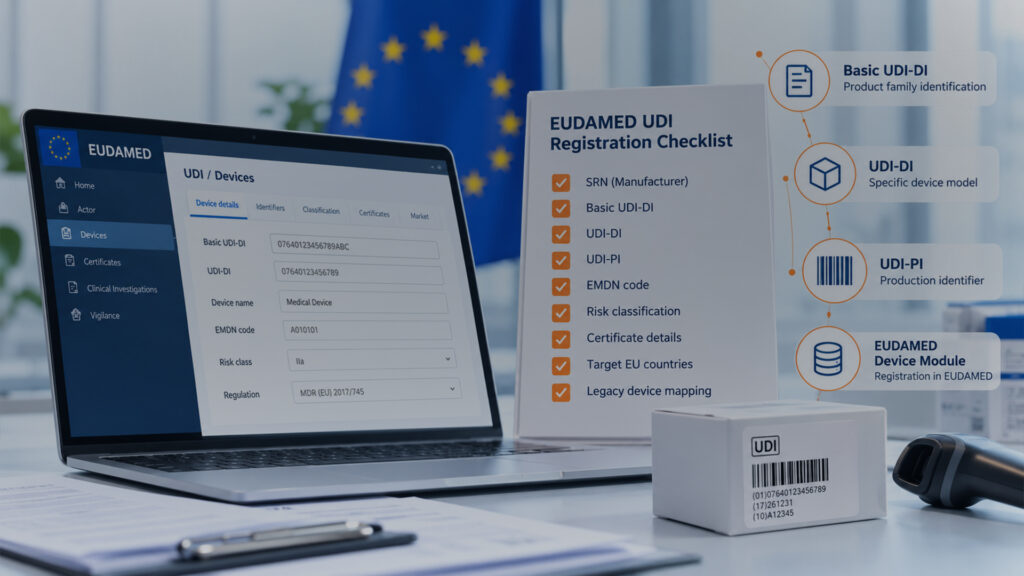
Manufacturers have been able to register in EUDAMED, the European database on medical devices, since December 2020 for the Actor
Using the EUDAMED UDI/Devices module became mandatory on 28 May 2026. This follows the Commission Decision (EU) 2025/2371, which formally
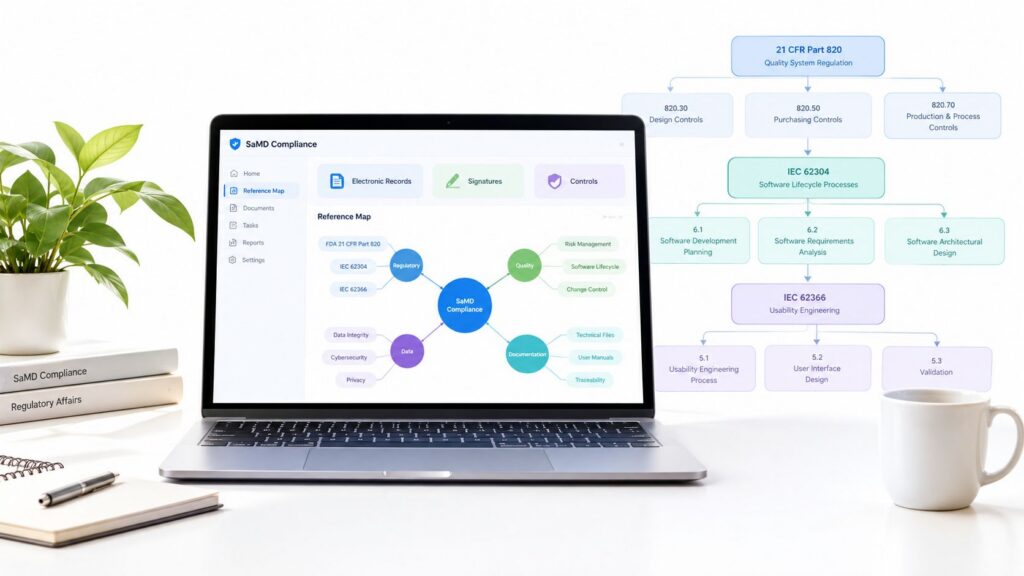
The Medical Device Regulation (EU) 2017/745 (MDR) has applied since 26 May 2021, yet software manufacturers are still working through
Explore AI-enabled SaMD requirements with our easy step-by-step guide.
Cookies help us improve your experience on our website. By using our site, you consent to the use of cookies as described in this policy.