The AI Act, Regulation (EU) 2024/1689, was adopted on 13 June 2024 and applies generally from 2 August 2026. Regulation
The investigational device exemption (IDE) is the framework that allows an unapproved device to be used in a clinical investigation
A Form FDA 483 and an FDA Warning Letter are not the same document. A 483 lists the investigator's observations
For medical devices, an FDA adverse event report is a Medical Device Report (MDR) under 21 CFR Part 803. The
A De Novo review is FDA's risk-based classification route for a novel device that has no legally marketed predicate, but
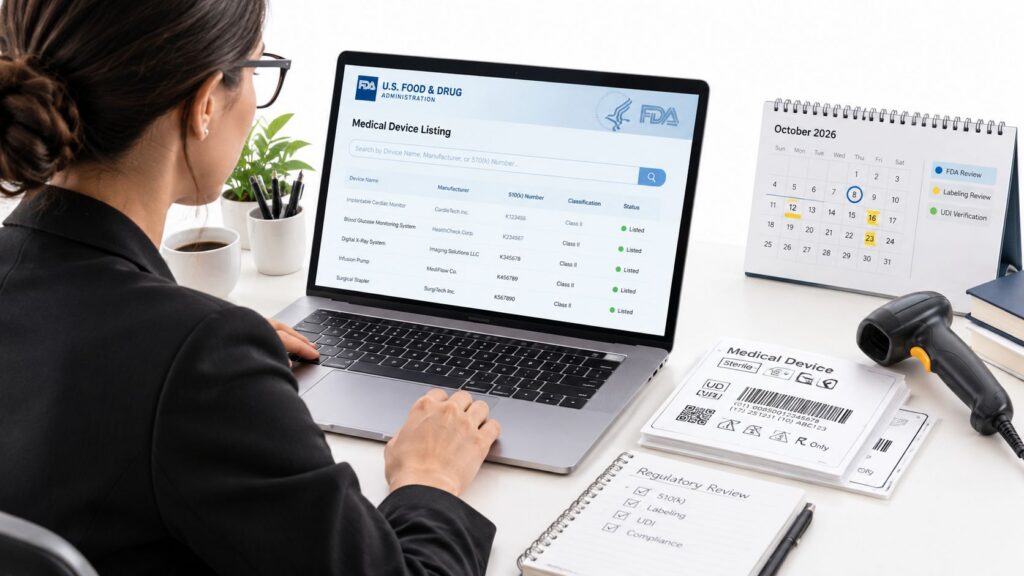
Device listing is the part of FDA registration that most teams touch once a year and otherwise forget. That is
ISO 27001 Requirements: A Clause-by-Clause Reference ISO/IEC 27001 has been the reference standard for information security management systems since 2005,
21 CFR Part 11 began as a permission, not a mandate. The FDA published it in March 1997, in 62
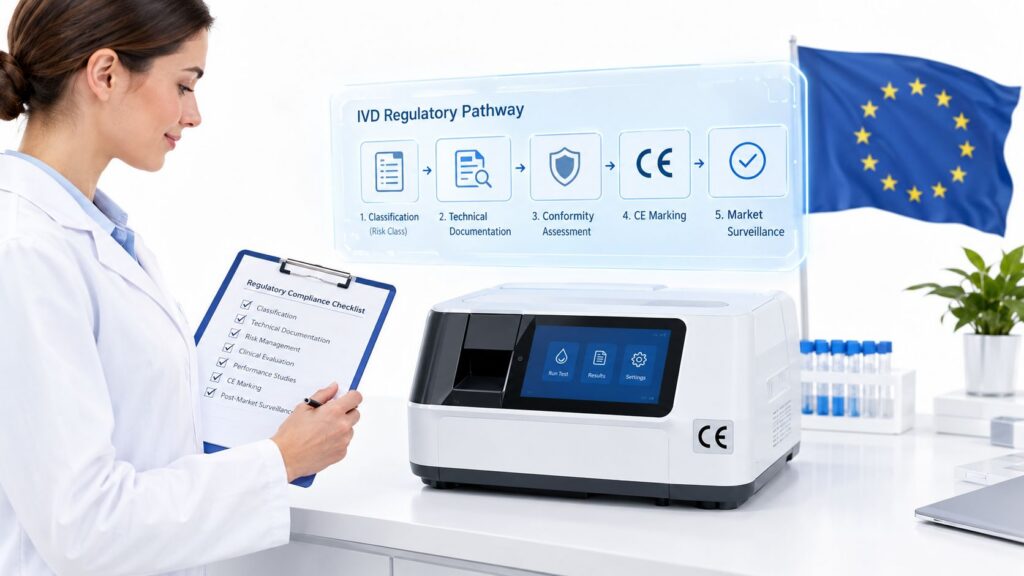
The shift from Directive 98/79/EC to Regulation (EU) 2017/746, the In Vitro Diagnostic Regulation, changed the route to IVD CE
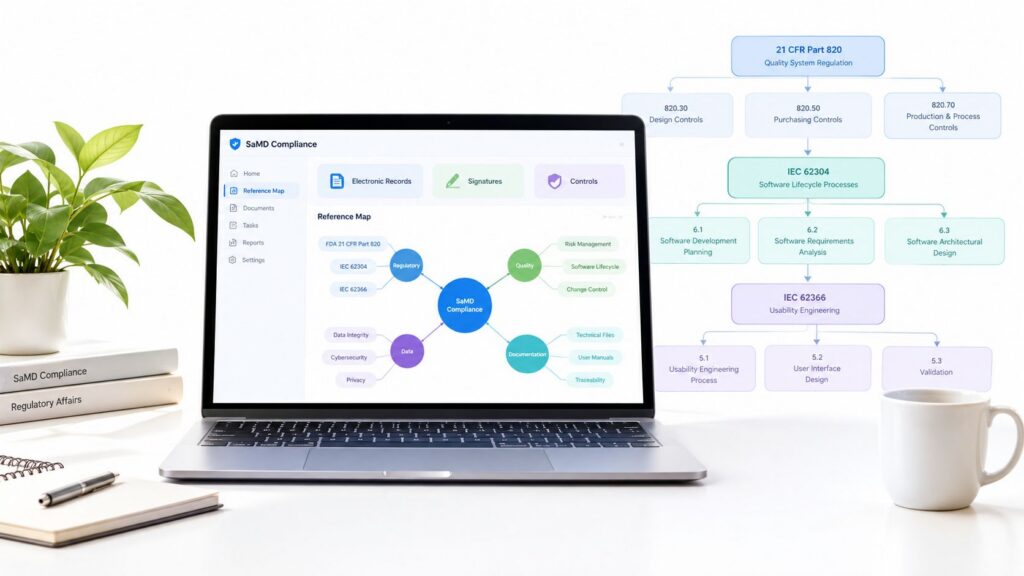
Explore AI-enabled SaMD requirements with our easy step-by-step guide.
Cookies help us improve your experience on our website. By using our site, you consent to the use of cookies as described in this policy.